Scientists Find How "Zombie" Cells Fuel Inflammation as We Age
A newly uncovered mitochondrial process explains why senescent "zombie" cells keep inflammation switched on long after it's needed — and blocking it improved tissue function and healthspan in mice.
Rashtriya Gaurav
6 min read · Sep 14, 2026
Key Takeaways
- Senescent "zombie" cells that build up with age stop dividing but stay metabolically active, secreting inflammatory molecules through a process called SASP.
- Researchers found that two mitochondrial signals converge to drive this: one exposes inflammatory genes, the other switches them on.
- Blocking the metabolic half of that process with a drug called CTPI-2 reduced inflammation across multiple tissues and improved healthspan in mice.
Inflammation is one of the immune system's most important defenses. When the body detects an infection or injury, it sends out alarm signals that recruit immune cells and other cells to eliminate the threat and begin repairing damaged tissue.
This response is essential, but it's supposed to shut down once the danger has passed. As people age, however, certain cells accumulate that can keep inflammation active for much longer than necessary. This persistent inflammation has been linked to numerous diseases that become more common later in life.
Researchers at Sanford Burnham Prebys Medical Discovery Institute, Mayo Clinic, and collaborating institutions have now uncovered a previously unknown connection between mitochondria, cell metabolism, and this chronic inflammation. Their findings, published in Nature, also showed that interfering with part of the process reduced inflammation and supported healthier aging in mice.
How "Zombie" Cells Accumulate With Age
Many cells in the body are capable of dividing, allowing us to grow and helping tissues repair themselves after injury. But as we get older, increasing numbers of cells enter a state known as senescence. These zombie-like senescent cells stop dividing, yet they remain alive and active.
"Senescent cells are not completely inert," said co-corresponding author Peter Adams, PhD, the Jeanne and Gary Herberger Leadership Chair in Cancer Research at Sanford Burnham Prebys. Adams also directs and is a professor in the Cancer Genome and Epigenetics Program. He explained that these cells stay metabolically active and run an inflammatory program that causes them to secrete inflammatory molecules.
When senescent cells switch on this inflammatory program, they're described as having the senescence-associated secretory phenotype, or SASP. SASP has been tied to the widespread inflammation associated with aging as well as a range of chronic diseases.
The research team, led by the laboratory of senior and co-corresponding author João Passos, PhD, a professor of Physiology at Mayo Clinic, set out to identify the molecular processes driving SASP, with a broader goal of finding ways to interrupt or reduce them.
Two separate mitochondrial signals turn out to work as partners: one pries open the DNA around inflammatory genes, the other flips the switch that turns them on.
Mitochondria Help Unlock Inflammatory Genes
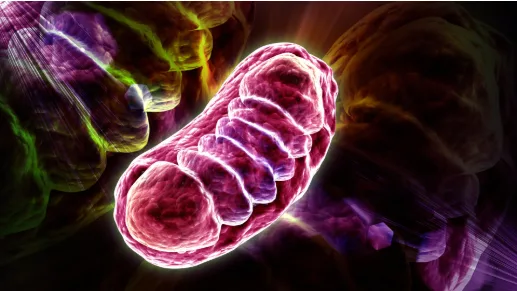
Although senescent cells no longer divide, they continue using energy and carrying out metabolic activity. The researchers found that their mitochondria — the structures responsible for much of a cell's energy production — behave differently from those in healthier cells, producing increased amounts of a molecule called acetyl-CoA.
Acetyl-CoA interacts with histones, the spool-like proteins around which DNA is wrapped. By altering these histones, acetyl-CoA can loosen how DNA is packaged. This doesn't change the genetic code itself — instead, it makes certain genes more accessible, including genes involved in the inflammatory SASP response. That metabolic change alone, however, wasn't enough to explain why senescent cells continuously release inflammatory molecules.
Two Mitochondrial Signals Work Together
A second process was also required. Damaged mitochondria can leak DNA and RNA into parts of the cell where those molecules normally shouldn't be present. The immune system interprets this misplaced genetic material as a danger signal, triggering an inflammatory response.
These immune signals activate transcription factors that help switch genes on. Those factors are then able to target the SASP genes that have already been made more accessible by elevated acetyl-CoA. In other words, two separate mitochondrial processes converge: one exposes inflammatory genes, while the other provides the immune signal needed to activate them.
"After seeing how these two independent pathways intersect, we wanted to see if interrupting one could prevent their partnership in promoting SASP," said Adams.
Blocking the Metabolic Signal Reduced Inflammation
To test that possibility, the researchers used a drug called CTPI-2, which blocks a transport protein responsible for carrying a component needed to produce acetyl-CoA. When CTPI-2 was tested in mice, it reduced inflammation throughout multiple tissues, and the treatment also improved tissue function and healthspan during aging.
Importantly, the immune signals caused by leaking mitochondrial DNA and RNA were still present. But by weakening the metabolic signal involving acetyl-CoA, the researchers made SASP genes more difficult to access and reduced their inflammatory effects.
"Even though the immune signaling from leaky mitochondria was still present, disrupting the metabolic signal made SASP genes less accessible and produced functional benefits," said Adams. He described using selective inhibitors like CTPI-2 to reduce acetyl-CoA, and in turn inflammation, as a novel therapeutic strategy worth exploring — and, more broadly, that targeting the metabolic signals controlling DNA accessibility could offer a new approach for easing age-related inflammation and functional decline.
The Bottom Line
Chronic, low-grade inflammation has long been recognized as a hallmark of aging, but the exact machinery keeping it switched on has been murky. This study offers a concrete mechanistic answer: two mitochondrial signals inside senescent cells team up to keep inflammatory genes both accessible and active. Interrupting just one half of that partnership was enough to calm inflammation and improve healthspan in mice — a promising, if still early-stage, lead for future therapies aimed at age-related disease.
This article is for informational purposes only and does not constitute medical advice. The findings described are based on animal and cell studies and have not been tested in human clinical trials.